
Last Updated: October 2026
What is eCTD 4.0, and when do you need to switch?
eCTD 4.0 is the ICH M8 successor to eCTD v3.2.2, the electronic format for submitting regulatory dossiers. It puts each submission into a single XML message, adds controlled vocabularies, and lets sponsors reuse previously submitted documents. The FDA has accepted new applications in v4.0 since September 16, 2024, without announcing an end date for v3.2.2. The EMA plans to make v4.0 mandatory for new centrally authorised marketing authorisation applications from Q1 2028.
In this article
- Where do FDA and EMA stand on eCTD 4.0?
- A quick history of eCTD
- What's new in eCTD 4.0?
- How does eCTD 4.0 affect your team?
- What are the benefits of eCTD 4.0?
- What do you need to do differently?
- What stays the same?
- Will eCTD 4.0 ever be fully implemented?
- How do you prepare for eCTD 4.0?
- How Kivo supports eCTD 4.0, and how the options compare
- FAQ
Where do FDA and EMA stand on eCTD 4.0?
Both agencies accept eCTD 4.0 today, and both still accept v3.2.2. Their timelines for requiring v4.0 differ, and the dates have moved before, so check each agency's own page before you plan around one.
| Agency | v4.0 accepted since | Scope today | Required from |
|---|---|---|---|
| FDA (CDER and CBER) | September 16, 2024 | New regulatory applications | No date announced; v3.2.2 still accepted |
| EMA | December 22, 2025 (optional) | New marketing authorisation applications for centrally authorised products | Q1 2028 for new centrally authorised MAAs, strongly recommended from Q1 2027 |
At the FDA, existing v3.2.2 applications cannot yet move to v4.0. The FDA says future phases will address forward compatibility for existing applications and two-way communication. Its current eCTD v4.0 Technical Conformance Guide is version 1.5, dated July 9, 2026.
In the EU, a pilot for new non-centrally authorised applications is planned for Q3/Q4 2026, and a forward-compatibility pilot for centrally authorised products is planned for Q4 2026. The EMA's updated eCTD v4.0 validation criteria apply from July 15, 2026. Japan's PMDA runs its own transition; check PMDA's current notices for its dates.
A quick history of eCTD
eCTD has been the standard format for electronic regulatory submissions for about two decades. Over that time the industry pushed the v3.2.2 XML standard to handle more submission types. Other formats, such as HL7's Regulated Product Submission (RPS), were discussed for drugs, biologics, devices, veterinary, and tobacco products. eCTD 4.0 is built on HL7 and RPS standards.
ICH began developing eCTD 4.0 around 2010 under its M8 working group and endorsed the implementation package, including controlled vocabularies, schemas, and transition guidance, in 2015. The new version extends XML functionality so documents can be reused and software can keep up with regulatory changes without a new DTD.
What's new in eCTD 4.0?
The biggest change is that one XML message now describes the whole submission, and documents are identified by unique IDs rather than by where they sit in a folder tree.
- Single XML backbone: Module 1 regional content and Modules 2 to 5 are described in one XML file.
- Document reuse: Unique identifiers (UUIDs) let you reference previously submitted content instead of resubmitting it.
- More lifecycle control: New lifecycle operators give finer control over how documents are updated.
- Controlled vocabularies (CVs): Standard metadata terms make classification and search consistent.
- Flatter structure: Placement comes from metadata, not a folder hierarchy.
How does eCTD 4.0 affect your team?
eCTD 4.0 changes how submissions are assembled and maintained more than how documents are written. The effects show up differently by role.
For executives
- Fewer software upgrades driven by DTD changes.
- Less submission rework through content reuse.
- Simpler multi-region submissions.
For regulatory affairs and operations
- Reviewers can see grouped, prioritised content.
- Fewer corrections caused by file-name errors, since metadata can be fixed directly.
- Faster assembly of multi-region applications.
What are the benefits of eCTD 4.0?
- Submit in the same format across ICH regions.
- Change document granularity while keeping lifecycle relationships.
- Set the order of documents within a CTD section.
- Flag content for additional processing, such as datasets.
- Reuse previously submitted documents.
- Use controlled vocabularies for consistent metadata.
- Correct attributes and metadata without resubmitting the document.
- Uncouple software upgrades from DTD updates.
What happens if you wait?
Nothing breaks today: the FDA and EMA both still accept v3.2.2. The risk is timing. For a new centrally authorised MAA in the EU, v4.0 is expected to be mandatory from Q1 2028, and teams that start late will be learning new metadata and tools while a filing is on the clock. Starting with education and metadata clean-up now is low cost.
What do you need to do differently for eCTD 4.0?
Your publishing software or publishing vendor handles most of the technical work. Your team's job is oversight, plus the metadata decisions that only you can make. The main new concepts are content reuse, lifecycle operators, context of use, and controlled vocabularies.
Reuse of content
UUIDs let you reuse content across sequences, regulatory activities, and applications. New sequences get smaller, and duplicate copies of the same document stop causing confusion.
The challenge for sponsors is tracking original content in their electronic document management system (EDMS) so links back to the originally submitted document are kept for lifecycle management.
New lifecycle operators
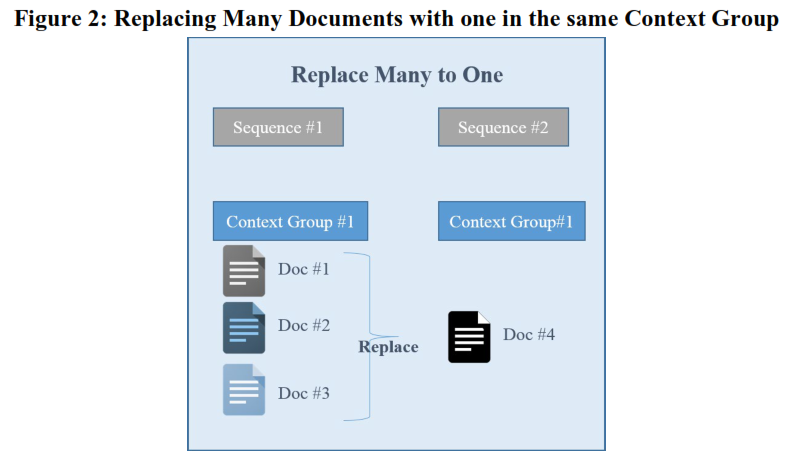
Lifecycle operators are now Active, Replace, and Suspend (previously New, Replace, and Delete). Replace can work one-to-one, one-to-many, or many-to-one while keeping lifecycle relationships on every document, and the change is tracked against the context of use.
FDA guidance gives this example:
Context of use
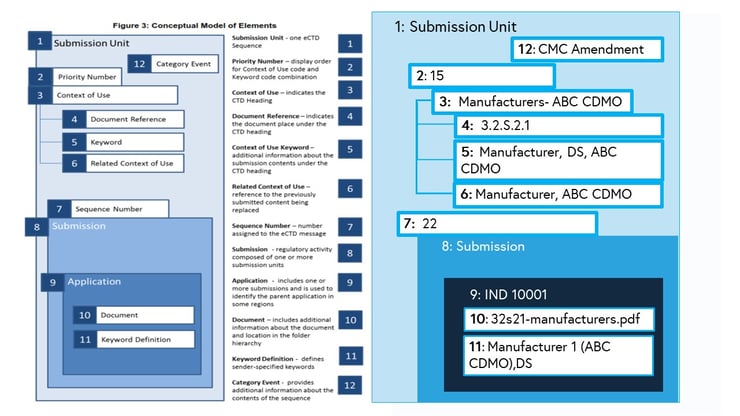
Context of use (CoU) plays a role similar to the leaf title in v3.2.2. It places a document in the CTD structure, carries its keywords, and records its lifecycle. Context of use, document type, and keywords together replace the study tagging file (STF) for organising study files in Modules 4 and 5, and they make the one-to-many and many-to-one replacements above possible.
Controlled vocabularies
Controlled vocabularies further describe content alongside the context of use. eCTD 4.0 accepts five kinds: ICH, HL7, regional, sender-defined, and external organisations such as ISO country codes.
Examples include ICH species terms such as mouse and rat, and US regional terms such as NDA, IND, and ANDA. New v4.0 vocabularies include Category Event, Document Type, and Study Group Order. Sponsors define their own keywords, such as a manufacturer's name (fictional example: ABC CDMO) or a drug name, and every region receives the same values because they travel in the XML.
ICH's implementation guide shows how context of use and keyword combinations place and describe Module 3 drug substance content:
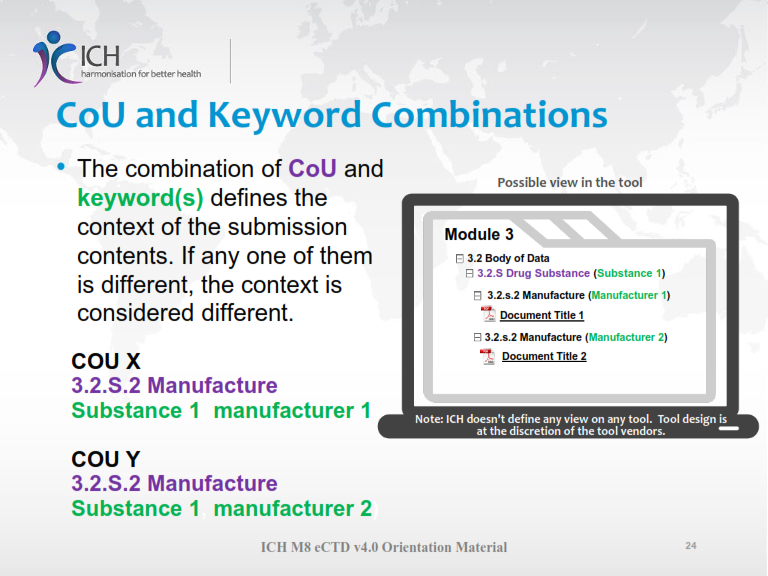
Below, a CMC amendment that adds a new contract manufacturer, ABC CDMO, for the drug substance is mapped to context of use, keywords, and the other required fields:
Metadata corrections
Typos in attributes and keywords can be corrected directly. Under v3.2.2, fixing a file name meant resubmitting the document.
Ordering and grouping documents
Priority numbers let you set the order of documents within a section, so reviewers see the most important content first. Grouping and ordering within a study organise Module 4 and 5 content and replace the STF.
No folder hierarchy
Files are delivered in a flat structure. Context of use and keywords place each file in the right location and add detail.
Viewing submissions
Older versions shipped a style sheet, so you could open the XML in a browser and navigate a sequence. v4.0 does not, so you need an eCTD viewer, either standalone or built into an EDMS, RIM, or publishing system.
What stays the same?
Authoring does not change: sponsors still write granular ICH CTD documents in Word for each section of the five modules and convert them to PDF.
Systems matter more, not less. Your EDMS has to track exactly which document version was submitted. Publishing tools must support v4.0 XML and every required vocabulary, including sender-defined ones. RIM systems often tie document management, submission tracking, and reporting together. For what to look for on the publishing side, see What eCTD Publishing and Validation Tools Should Include.
Will eCTD 4.0 ever be fully implemented?
Possibly not in the way v3.2.2 was. eCTD 4.0 is still a way of packaging documents and sending them to an agency, and some regulators and sponsors are already working on exchanging structured data in near real time instead. If that model matures, it could take over parts of the job before every region makes v4.0 mandatory.
Full adoption is already slow. More than two years after the FDA began accepting v4.0, it has not set a date to require it or to move existing v3.2.2 applications across, and the EMA's mandatory date applies only to new centrally authorised applications. Meanwhile, two efforts point past document-based submissions:
- Structured quality data at the FDA. The FDA's Pharmaceutical Quality/Chemistry, Manufacturing and Controls (PQ/CMC) project is defining structured data for parts of Module 3, using HL7 FHIR as the exchange format. The first part of the FHIR implementation guide has been published as a Standard for Trial Use. The FDA says the standard is not meant to structure all eCTD quality information, and it has not announced when it will accept or require it. Its companion KASA system (Knowledge-aided Assessment and Structured Application) is built to assess that structured data.
- Cloud-based exchange between sponsors and regulators. Accumulus Synergy, a nonprofit industry association backed by large biopharma companies, launched a platform in February 2024 for what it calls real-time information exchange between sponsors and the agencies that review their products. The association reports that regulators in pilots have used it to ask questions and receive sponsor answers on post-approval change submissions, and that one post-approval CMC dossier was delivered to a reference country and nearly two dozen reliance countries at the same time. These are the association's own reports, and participation so far centres on pilots and reliance procedures rather than primary approvals.
Neither effort replaces eCTD today, and neither has a date when it would. For now, the practical reading is that eCTD 4.0 is the format to plan for over the next few years. The longer-term direction is data that is structured once and reused, which is the same thing that makes v4.0 work: consistent metadata, controlled vocabularies, and a single source of truth for each document. Work on those now and it pays off whichever model wins.
How do you prepare for eCTD 4.0?
Start with education and metadata, then line up your tools before an agency deadline forces the issue.
- Train your team on context of use, controlled vocabularies, and lifecycle operators.
- Build a sponsor keyword list and align it with the metadata in your EDMS and RIM system.
- Ask your publishing vendor or tool provider for its v4.0 support and validation status, for each agency you file with.
- Decide which new application should be your first v4.0 filing. At the FDA, new applications can be filed in v4.0 now; in the EU, contact the EMA eCTD v4.0 team before your first submission.
- Check each agency's eCTD v4.0 page before you lock a plan, since timelines are still being updated.
How Kivo supports eCTD 4.0, and how the options compare
Kivo RIM includes an eCTD viewer that supports eCTD 4.0. It is browser-based, works on Mac and PC, and links live to the Kivo document management system, so each submitted document's metadata and audit trail are one click away. Kivo also provides 450+ pre-formatted ICH templates, including ICH M11 clinical templates, and tracks submissions, correspondence, and commitments in one place. Kivo does not publish eCTDs itself: it prepares submission-ready packages for your chosen publishing partner and keeps submitted eCTDs viewable and linked to their source documents. Kivo is rated 4.9/5 on G2 and is used by 200+ sponsor, consultant, and CRO teams.
Several vendors offer eCTD viewing, archiving, or publishing, usually inside a broader regulatory suite. The table below summarises how Kivo and three alternatives handle eCTD, based on each vendor's published materials.
| Option | Product area | eCTD handling (per vendor materials) | Typically fits |
|---|---|---|---|
| Kivo | Kivo RIM eCTD Viewer, linked to Kivo DMS | Browser-based viewer with sequence, current, and cumulative views; live links to DMS metadata and audit trails; supports eCTD 4.0. Publishing is handled by a partner. | Clinical-stage teams that want viewing tied to document management and RIM |
| Veeva | Vault Submissions Archive | Cloud viewer for eCTD, NeeS, and legacy formats; current, sequential, and regulatory-action views; preserves the eCTD XML backbone and hyperlinks on import. | Teams already on the Veeva Vault platform |
| Ennov | Ennov Regulatory Suite (Ennov Dossier) | Generates, manages, publishes, validates, and archives eCTD, NeeS, and eCopy submissions within a suite that includes document management and RIM. | Teams that want publishing and RIM in one suite |
| ArisGlobal | LifeSphere Publishing / Regulatory | Plans, compiles, publishes, and validates submissions with eCTD lifecycle management, integrated with document management for review. | Teams that want publishing and RIM on one platform |
This table is not an independent ranking. It does not assess implementation quality, market share, or fit for any particular organisation, and capabilities change, so confirm details, including eCTD 4.0 support, with each vendor and validate against your own requirements.
FAQ
Is eCTD 4.0 mandatory?
Not yet at the FDA, which accepts v4.0 for new applications but has not announced a date to stop accepting v3.2.2. In the EU, the EMA plans to require v4.0 for new centrally authorised marketing authorisation applications from Q1 2028, with use strongly recommended from Q1 2027.
Can I move an existing application from v3.2.2 to eCTD 4.0?
Not at the FDA yet. Its current phase covers new applications only, and forward compatibility for existing v3.2.2 applications is planned for a later phase. The EMA plans a forward-compatibility pilot for centrally authorised products starting in Q4 2026.
What is context of use in eCTD 4.0?
Context of use is the element that places a document in the CTD structure, carries its keywords, and tracks its lifecycle. It takes over much of the role the leaf title and folder location played in v3.2.2, and it enables one-to-many and many-to-one document replacement.
Do I need new software for eCTD 4.0?
Usually your publishing tool or vendor needs to support v4.0, and you need a viewer that can open v4.0 sequences, since the format no longer ships a browser style sheet. Your document management system also needs metadata that maps to the new controlled vocabularies.
What stays the same with eCTD 4.0?
Document authoring. You still write granular ICH CTD documents in Word for each module and convert them to PDF. What changes is how those documents are described, placed, and reused in the submission's XML.
References
- FDA: Electronic Common Technical Document (eCTD) v4.0
- FDA: eCTD Submission Standards for eCTD v4.0 and Regional M1
- EMA eSubmission: eCTD v4.0
- FDA: Pharmaceutical Quality/Chemistry, Manufacturing & Controls (PQ/CMC)
- Accumulus Synergy: platform launch announcement (February 2024)
- ICH eCTD v4.0 implementation package
